Sailgene Technology
 WhatsApp
 service@sailgene.com
Introduction
A haplotype-resolved telomere-to-telomere (T2T) genome aims to reconstruct each haplotype independently and generate complete chromosome sequences extending from one telomere to the other.
Unlike conventional genome assemblies that may collapse homologous chromosomes into a single consensus sequence, haplotype-resolved T2T assembly preserves the sequence and structural differences between haplotypes while maximizing chromosome completeness.
The resulting genome provides a more comprehensive representation of genetic variation, including heterozygous regions, structural differences, repetitive sequences, centromeres, telomeres, and other complex genomic regions that are often difficult to resolve in conventional assemblies.
Applications

Heterozygous and Hybrid Genome Research

Structural and Haplotype Variation Analysis

Evolutionary and Functional Genomics

Pangenome and Molecular Breeding
Highlights

Haplotype-Resolved Genome Reconstruction

Telomere-to-Telomere Completeness

Resolution of Complex Genomic Regions

Comprehensive Genomic Resource for Advanced Research
Workflow


Library QC


Bioinformatics Ananlysis
Haplotype-resolved T2T Demo Report
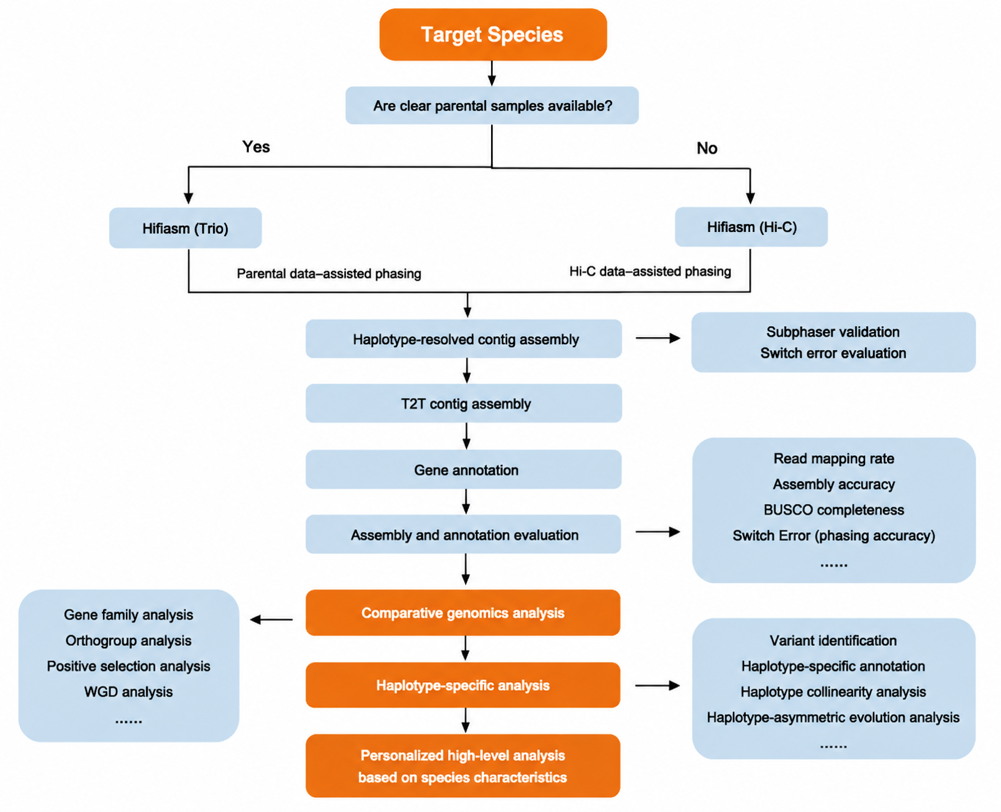
Analysis workflow
Publications
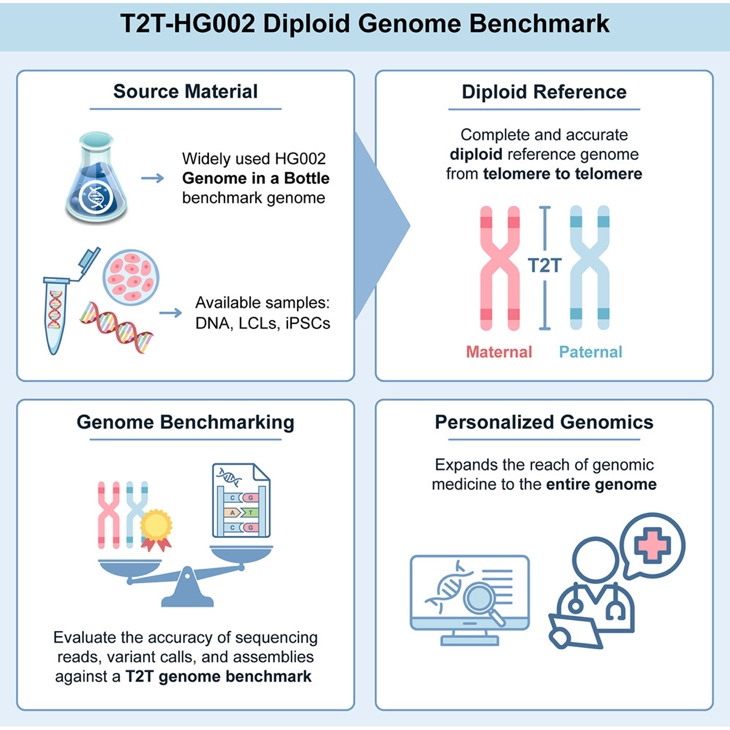
A complete diploid human genome benchmark for personalized genomics
Journal: Cell
IF: 45.1 (2026)
Highlights
• A telomere-to-telomere diploid assembly of HG002 achieves near-perfect accuracy
• Personalized diploid genome annotation reveals haplotype-specific gene variation
• Companion software evaluates sequence, assembly, and variant accuracy genome-wide
• New benchmark facilitates transition to genome inference and personalized genomics
Summary
Human genome sequencing typically relies on mapping reads to a reference genome to call variants, but this approach introduces technical biases, excluding duplicated and structurally polymorphic regions of the genome. To overcome this, we present a telomere-to-telomere genome benchmark with near-perfect accuracy across 99.4% of the diploid HG002 genome. This benchmark adds 701.4 Mb of autosomal sequence and both sex chromosomes (216.8 Mb), which were absent from prior benchmarks. We annotated genes and repeats on both haplotypes, including 19,956 protein-coding genes on the maternal haplotype and 19,190 on the paternal haplotype, and developed new methods to measure the accuracy of reads, phased variant call sets, and assemblies against a diploid reference. Genome-wide analyses show that de novo assembly resolves 2%–7% more sequence and outperforms variant calling accuracy by an order of magnitude, expanding the reach of genomic medicine to the entire genome and enabling a new era of personalized genomics.
Contact Us
If you are interested in our long-read sequencing services or potential collaboration, please contact us. Our team is ready to support your research with tailored solutions. We also welcome feedback from users to help us improve our services.
Contact Us
Add: One Innovation Drive, Suite B3-406, Worcester, MA 01605, USA
在线客服添加返回顶部
右侧在线客服样式 1,2,3 1
图片alt标题设置: SAILGENE TECHNOLOGY INC.
表单验证提示文本: Content cannot be empty!
循环体没有内容时: Sorry,no matching items were found.
CSS / JS 文件放置地